Neural-network based simulation of rare event processes at the water/oxide interface
Subproject P12
Atomistic computer simulations of processes occurring at the water/oxide interface are challenging in several ways. The calculation of atomic forces based on ab initio methods is computationally very demanding, and barrier crossing events may lead to long computation times. Both these aspects severely limit accessible system sizes and simulation times.
Building on the neural network potentials and the rare events simulation methods developed in the first funding period, project P12 will simulate complex dynamical processes occurring at the oxide/water interface. In particular, one central objective will be to investigate heterogeneous ice nucleation on various mineral surfaces that are of atmospheric significance. The studies will involve tight interactions with projects P03 Kresse and P02 Diebold. In addition, project P12 will continue to explore the use of machine learning approaches for trajectory-based rare events sampling. Here, the main objectives are to develop efficient latent space methods to sample path distributions and the on-the-fly optimization of transition path sampling simulations based on information encoded in learned committor functions.
Christoph Dellago
PI
Expertise
Our research efforts focus on the development of simulation algorithms and their application to investigate dynamical processes in condensed matter systems based on the principles of equilibrium and non-equilibrium statistical mechanics. In particular, we have helped to create the transition path sampling methodology for the simulation of rare but important events, such as nucleation aprocesses, chemical reactions and biomolecular reorganizations. More recently, we have worked on applying machine learning methods to molecular structure recognition and the representation of potential and free energy surfaces.
Recent research topics include:
- Self-assembly of nanocrystals
- Folding and unfolding of biopolymers
- Interfaces in aqueous systems
- Phase separation in alloys
- Thermo-polarisation
- Structure and dynamics of water and ice
- Cavitation
- Crystallization
- Non-equilibrium work fluctuations
Team
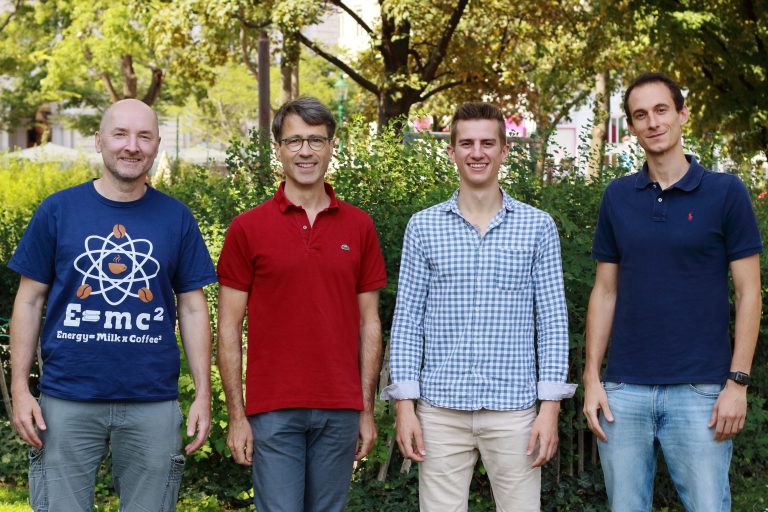
Christoph Dellago
PI
Pablo Montero de Hijes
PostDoc

Alexander Gorfer
PhD Student
Salvatore Romano
PhD Student

Justin Pils
PhD Student

Marco Janach
PhD Student
Associates
Alessandro Coretti
PostDoc
Sebastian Falkner
PhD Student

Maximilian Negedly
PhD Student
Publications
2026
de Hijes, Pablo Montero; Neubeck, L.; Kresse, Georg; Dellago, Christoph
Dynamical properties of ab initio water from machine-learning potentials Journal Article
In: The Journal of Chemical Physics, vol. 165, pp. 074502, 2026, (2026 JCP Emerging Investigators Special Collection).
Abstract | Links | BibTeX | Tags: P03, P12
@article{Montero_2026c,

title = {Dynamical properties of ab initio water from machine-learning potentials},

author = {Pablo Montero de Hijes and L. Neubeck and Georg Kresse and Christoph Dellago},

doi = {10.1063/5.0348453},

year  = {2026},

date = {2026-08-17},

journal = {The Journal of Chemical Physics},

volume = {165},

pages = {074502},

abstract = {We assess the dynamical properties of liquid water predicted by several density functionals using machine-learning interatomic potentials. MACE models were trained for SCAN, RPBE-D3/zd, revPBE-D3/zd, revPBE0-D3/BJ, PBE0-D3/zd, and PBE0-D3/BJ using previously reported \emph{ab initio} datasets. We compare translational, rotational, and viscous dynamics through time-correlation functions, which resolve relaxation processes across different timescales, and through the corresponding long-time kinetic coefficients. The diffusion coefficient, second-rank orientational relaxation time, and shear viscosity reveal systematic differences among functionals. Part of these differences can be rationalized as shifts along the phase diagram, as comparisons relative to the melting temperature of each functional reduce the spread in the dynamical observables. Among the functionals considered, RPBE-D3/zd provides the best overall agreement with experiment. We, therefore, perform a broader validation of RPBE-D3/zd using a Behler–Parrinello neural-network potential over a wide range of temperatures, densities, and pressures. The model reproduces the magnitude and anomalous pressure dependence of the diffusion coefficient, gives generally good viscosities, and captures the temperature dependence of the rotational relaxation time.},

note = {2026 JCP Emerging Investigators Special Collection},

keywords = {P03, P12},

pubstate = {published},

tppubtype = {article}

}
Falkner, Sebastian; Schwierz, Nadine; de Hijes, Pablo Montero
Unbiased exploration of the transition region in ice nucleation using the NpH ensemble Journal Article
In: The Journal of Chemical Physics, vol. 165, pp. 054504, 2026, (Festschrift in honor of Christoph Dellago).
Abstract | Links | BibTeX | Tags: P12
@article{Falkner_2026a,

title = {Unbiased exploration of the transition region in ice nucleation using the NpH ensemble},

author = {Sebastian Falkner and Nadine Schwierz and Pablo Montero de Hijes},

doi = {10.1063/5.0336532},

year  = {2026},

date = {2026-08-05},

urldate = {2026-08-05},

journal = {The Journal of Chemical Physics},

volume = {165},

pages = {054504},

abstract = {In this work, we employ the isenthalpic–isobaric (NpH) ensemble to sample the transition region of ice nucleation without any external bias, thereby avoiding potentially artificial memory effects introduced by projections onto collective variables. Within this framework, we identify relevant degrees of freedom that expose the intrinsically non-Markovian nature of the largest nucleus size, indicating that it is not sufficient on its own to describe nucleation dynamics. The NpH ensemble leads to long-lived nuclei through the coupling between latent heat release or absorption and temperature fluctuations. As a result, nuclei persist over extended timescales and undergo a slow internal evolution, which we refer to as aging. A signature of this behavior is the emergence of hysteresis in the largest cluster size–temperature plane. To quantify these effects, we perform a structural analysis based on a high-dimensional set of descriptors, which we project onto a low-dimensional latent space using a neural network-based autoencoder. This approach reveals the existence of structurally distinct classes of nuclei with similar sizes and temperatures. Finally, we compare the nucleus sizes obtained with this approach with those from previous studies employing different methodologies, finding good agreement.},

note = {Festschrift in honor of Christoph Dellago},

keywords = {P12},

pubstate = {published},

tppubtype = {article}

}
de Hijes, Pablo Montero; Falkner, Sebastian; Dellago, Christoph
Non-Markovian dynamics in ice nucleation Journal Article
In: The Journal of Chemical Physics, vol. 164, iss. 9, pp. 094501, 2026.
Abstract | Links | BibTeX | Tags: P12
@article{Montero_2026b,

title = {Non-Markovian dynamics in ice nucleation},

author = {Pablo Montero de Hijes and Sebastian Falkner and Christoph Dellago},

doi = {10.1063/5.0314412},

year  = {2026},

date = {2026-03-02},

journal = {The Journal of Chemical Physics},

volume = {164},

issue = {9},

pages = {094501},

abstract = {In simulation studies of crystallization, the size of the largest crystalline nucleus is often used as a reaction coordinate to monitor the progress of the nucleation process. Here, we investigate, for the case of homogeneous ice nucleation, whether the nucleus size exhibits Markovian dynamics, as assumed in classical nucleation theory. Using 300 independent nucleation trajectories generated by molecular dynamics, we evaluate the mean recurrence time required to reach selected values of the largest nucleus size. Early recurrences consistently take longer than later ones, revealing a clear history dependence and thus non-Markovian dynamics. To identify the slow modes underlying this behavior, we analyze several structural descriptors of the nucleus, observing subtle but systematic differences between nuclei at early and late recurrences. By training a neural network on 2700 short trajectories to learn the committor, we identify relevant collective variables. Based on these features, symbolic regression provides a compact approximation of the committor, that is, an improved reaction coordinate, which we subsequently test for Markovian dynamics.},

keywords = {P12},

pubstate = {published},

tppubtype = {article}

}
de Hijes, Pablo Montero; Shi, K; Vega, C; Dellago, Christoph
Comparing the Mechanical and Thermodynamic Definitions of Pressure in Ice Nucleation Journal Article
In: The Journal of Physical Chemistry Letters, 2026.
Abstract | Links | BibTeX | Tags: P12
@article{Montero_2026a,

title = {Comparing the Mechanical and Thermodynamic Definitions of Pressure in Ice Nucleation},

author = {Pablo Montero de Hijes and K Shi and C Vega and Christoph Dellago},

doi = {10.1021/acs.jpclett.5c03700},

year  = {2026},

date = {2026-02-12},

journal = {The Journal of Physical Chemistry Letters},

abstract = {Crystal nucleation studies using hard-sphere and Lennard-Jones models have shown that the actual (mechanical) pressure within the nucleus is lower than that in the surrounding liquid. Here, we use the mechanical route to obtain the pressure for an ice nucleus in supercooled water (TIP4P/Ice) at 1 bar and 247 K. From this pressure, we obtain the interfacial stress using a thermodynamic definition consistent with mechanical arguments. Moreover, we compare the mechanical pressure with the thermodynamic pressure of bulk ice at an equal chemical potential and the interfacial stress with the interfacial free energy. Furthermore, we investigate these properties on the basal plane. We find that unlike in hard-sphere and Lennard-Jones systems, mechanical and thermodynamic pressures agree for the nucleus, and the interfacial stress and free energy are comparable. However, the basal interface displays an interfacial stress nearly twice its interfacial free energy, suggesting that this agreement may be dependent on the system, underscoring the limitations of mechanical routes to solid–liquid interfacial free energies.},

keywords = {P12},

pubstate = {published},

tppubtype = {article}

}
2025
Coretti, Alessandro; Falkner, Sebastian; Geissler, Phillip; Dellago, Christoph
Learning Mappings between Equilibrium States of Liquid Systems Using Normalizing Flows Journal Article
In: The Journal of Chemical Physics, vol. 162, iss. 18, pp. 184102, 2025.
Abstract | Links | BibTeX | Tags: P12
@article{Coretti2022,

title = {Learning Mappings between Equilibrium States of Liquid Systems Using Normalizing Flows},

author = {Alessandro Coretti and Sebastian Falkner and Phillip Geissler and Christoph Dellago},

doi = {10.1063/5.0253034},

year  = {2025},

date = {2025-05-08},

urldate = {2022-08-22},

journal = {The Journal of Chemical Physics},

volume = {162},

issue = {18},

pages = {184102},

abstract = {Generative models and, in particular, normalizing flows are a promising tool in statistical mechanics to address the sampling problem in condensed-matter systems. In this work, we investigate the potential of normalizing flows to learn a transformation to map different liquid systems into each other while allowing at the same time to obtain an unbiased equilibrium distribution. We apply this methodology to the mapping of a small system of fully repulsive disks modeled via the Weeks–Chandler–Andersen potential into a Lennard-Jones system in the liquid phase at different coordinates in the phase diagram. We obtain an improvement in the relative effective sample size of the generated distribution up to a factor of six compared to direct reweighting. We show that this factor can have a strong dependency on the thermodynamic parameters of the source and target system.},

keywords = {P12},

pubstate = {published},

tppubtype = {article}

}
Gorfer, Alexander; Abart, Rainer; Dellago, Christoph
Mechanism and kinetics of sodium diffusion in Na-feldspar from neural network based atomistic simulations Journal Article
In: Acta Materialia, vol. 286, pp. 120657, 2025, (Acta Materialia, submitted).
Abstract | Links | BibTeX | Tags: P12
@article{Gorfer_2024b,

title = {Mechanism and kinetics of sodium diffusion in Na-feldspar from neural network based atomistic simulations},

author = {Alexander Gorfer and Rainer Abart and Christoph Dellago},

doi = {10.1016/j.actamat.2024.120657},

year  = {2025},

date = {2025-03-01},

urldate = {2024-05-29},

journal = {Acta Materialia},

volume = {286},

pages = {120657},

abstract = {Alkali diffusion is a first-order control for microstructure and compositional evolution of feldspar during cooling from high temperatures of primary magmatic or metamorphic crystallization, and knowledge of the respective diffusion coefficients is crucial for reconstructing thermal histories. Our understanding of alkali diffusion in feldspar is, however, hindered by an insufficient grasp of the underlying diffusion mechanisms. We performed molecular dynamics simulations of sodium feldspar (Albite) containing different point defects using a recently developed neural network potential. A high degree of agreement between the sodium self-diffusion coefficients obtained from model simulations and those determined experimentally in earlier studies motivated a detailed investigation into the interstitial and vacancy mechanisms, corresponding jump rates, correlation factors and anisotropy. We identified a dumbbell shaped double occupancy of an alkali site as an important point defect and a correlation effect originating from the orientation of the dumbbell as a possible cause for the ⊥(001)>⊥(010) diffusion anisotropy, which has been reported in a slew of feldspar cation diffusion experiments.},

note = {Acta Materialia, submitted},

keywords = {P12},

pubstate = {published},

tppubtype = {article}

}
Kývala, Lukáš; Hijes, Pablo Montero De; Dellago, Christoph
Unsupervised identification of crystal defects from atomistic potential descriptors Journal Article
In: npj Computational Materials, vol. 11, iss. 1, pp. 50, 2025.
Abstract | Links | BibTeX | Tags: P12
@article{Kyvala_2024a,

title = {Unsupervised identification of crystal defects from atomistic potential descriptors},

author = {Lukáš Kývala and Pablo Montero De Hijes and Christoph Dellago},

doi = {10.1038/s41524-025-01544-2},

year  = {2025},

date = {2025-02-27},

urldate = {2024-05-02},

journal = {npj Computational Materials},

volume = {11},

issue = {1},

pages = {50},

abstract = {Identifying crystal defects is vital for unraveling the origins of many physical phenomena. Traditionally used order parameters are system-dependent and can be computationally expensive to calculate for long molecular dynamics simulations. Unsupervised algorithms offer an alternative independent of the studied system and can utilize precalculated atomistic potential descriptors from molecular dynamics simulations. We compare the performance of three such algorithms (PCA, UMAP, and PaCMAP) on silicon and water systems. Initially, we evaluate the algorithms for recognizing phases, including crystal polymorphs and the melt, followed by an extension of our analysis to identify interstitials, vacancies, and interfaces. While PCA is found unsuitable for effective classification, it has been shown to be a suitable initialization for UMAP and PaCMAP. Both UMAP and PaCMAP show promising results overall, with PaCMAP proving more robust in classification, except in cases of significant class imbalance, where UMAP performs better. Notably, both algorithms successfully identify nuclei in supercooled water, demonstrating their applicability to ice nucleation in water.},

keywords = {P12},

pubstate = {published},

tppubtype = {article}

}

Romano, Salvatore; de Hijes, Pablo Montero; Meier, Matthias; Kresse, Georg; Franchini, Cesare; Dellago, Christoph
Structure and Dynamics of the Magnetite(001)/Water Interface from Molecular Dynamics Simulations Based on a Neural Network Potential Journal Article
In: Journal of Chemical Theory and Computation, vol. 21, iss. 4, pp. 1951–1960, 2025.
Abstract | Links | BibTeX | Tags: P03, P07, P12
@article{Romano_2024a,

title = {Structure and Dynamics of the Magnetite(001)/Water Interface from Molecular Dynamics Simulations Based on a Neural Network Potential},

author = {Salvatore Romano and Pablo Montero de Hijes and Matthias Meier and Georg Kresse and Cesare Franchini and Christoph Dellago},

doi = {10.1021/acs.jctc.4c01507},

year  = {2025},

date = {2025-02-13},

urldate = {2024-08-21},

journal = {Journal of Chemical Theory and Computation},

volume = {21},

issue = {4},

pages = {1951–1960},

abstract = {The magnetite/water interface is commonly found in nature and plays a crucial role in various technological applications. However, our understanding of its structural and dynamical properties at the molecular scale remains still limited. In this study, we developed an efficient Behler-Parrinello neural network potential (NNP) for the magnetite/water system, paying particular attention to the accurate generation of reference data with density functional theory. Using this NNP, we performed extensive molecular dynamics simulations of the magnetite (001) surface across a wide range of water coverages, from single molecules to bulk water. Our simulations revealed several new ground states of low coverage water on the Subsurface Cation Vacancy (SCV) model and yielded a density profile of water at the surface that exhibits marked layering. By calculating mean square displacements, we obtained quantitative information on the diffusion of water molecules on the SCV for different coverages, revealing significant anisotropy. Additionally, our simulations provided qualitative insights into the dissociation mechanisms of water molecules at the surface.},

keywords = {P03, P07, P12},

pubstate = {published},

tppubtype = {article}

}
2024

Romano, Salvatore; Kaur, Harsharan; Zelenka, Moritz; Hijes, Pablo Montero De; Eder, Moritz; Parkinson, Gareth S.; Backus, Ellen H. G.; Dellago, Christoph
Structure of the water/magnetite interface from sum frequency generation experiments and neural network based molecular dynamics simulations Journal Article
In: arXiv, 2024.
Abstract | Links | BibTeX | Tags: P04, P11, P12
@article{Romano_2024b,

title = {Structure of the water/magnetite interface from sum frequency generation experiments and neural network based molecular dynamics simulations},

author = {Salvatore Romano and Harsharan Kaur and Moritz Zelenka and Pablo Montero De Hijes and Moritz Eder and Gareth S. Parkinson and Ellen H. G. Backus and Christoph Dellago},

url = {https://arxiv.org/abs/2410.12717},

year  = {2024},

date = {2024-10-16},

urldate = {2024-10-16},

journal = {arXiv},

abstract = {Magnetite, a naturally abundant mineral, frequently interacts with water in both natural settings and various technical applications, making the study of its surface chemistry highly relevant. In this work, we investigate the hydrogen bonding dynamics and the presence of hydroxyl species at the magnetite-water interface using a combination of neural network potential-based molecular dynamics simulations and sum frequency generation vibrational spectroscopy. Our simulations, which involved large water systems, allowed us to identify distinct interfacial species, such as dissociated hydrogen and hydroxide ions formed by water dissociation. Notably, water molecules near the interface exhibited a preference for dipole orientation towards the surface, with bulk-like water behavior only re-emerging beyond 60 Å from the surface. The vibrational spectroscopy results aligned well with the simulations, confirming the presence of a hydrogen bond network in the surface ad-layers. The analysis revealed that surface-adsorbed hydroxyl groups orient their hydrogen atoms towards the water bulk. In contrast, hydrogen-bonded water molecules align with their hydrogen atoms pointing towards the magnetite surface.},

keywords = {P04, P11, P12},

pubstate = {published},

tppubtype = {article}

}
de Hijes, Pablo Montero; Dellago, Christoph; Jinnouchi, Ryosuke; Kresse, Georg
Density isobar of water and melting temperature of ice: Assessing common density functionals Journal Article
In: The Journal of Chemical Physics, vol. 161, pp. 131102, 2024.
Abstract | Links | BibTeX | Tags: P03, P12
@article{Montero-de-Hijes_2024b,

title = {Density isobar of water and melting temperature of ice: Assessing common density functionals},

author = {Pablo Montero de Hijes and Christoph Dellago and Ryosuke Jinnouchi and Georg Kresse},

url = {https://doi.org/10.1063/5.0227514},

year  = {2024},

date = {2024-10-03},

urldate = {2024-06-06},

journal = {The Journal of Chemical Physics},

volume = {161},

pages = {131102},

abstract = {We investigate the density isobar of water and the melting temperature of ice using six different density functionals. Machine-learning potentials are employed to ensure computational affordability. Our findings reveal significant discrepancies between various base functionals. Notably, even the choice of damping can result in substantial differences. Overall, the outcomes obtained through density functional theory are not entirely satisfactory across most utilized functionals. All functionals exhibit significant deviations either in the melting temperature or equilibrium volume, with most of them even predicting an incorrect volume difference between ice and water. Our heuristic analysis indicates that a hybrid functional with 25% exact exchange and van der Waals damping averaged between zero and Becke–Johnson dampings yields the closest agreement with experimental data. This study underscores the necessity for further enhancements in the treatment of van der Waals interactions and, more broadly, density functional theory to enable accurate quantitative predictions for molecular liquids.},

keywords = {P03, P12},

pubstate = {published},

tppubtype = {article}

}